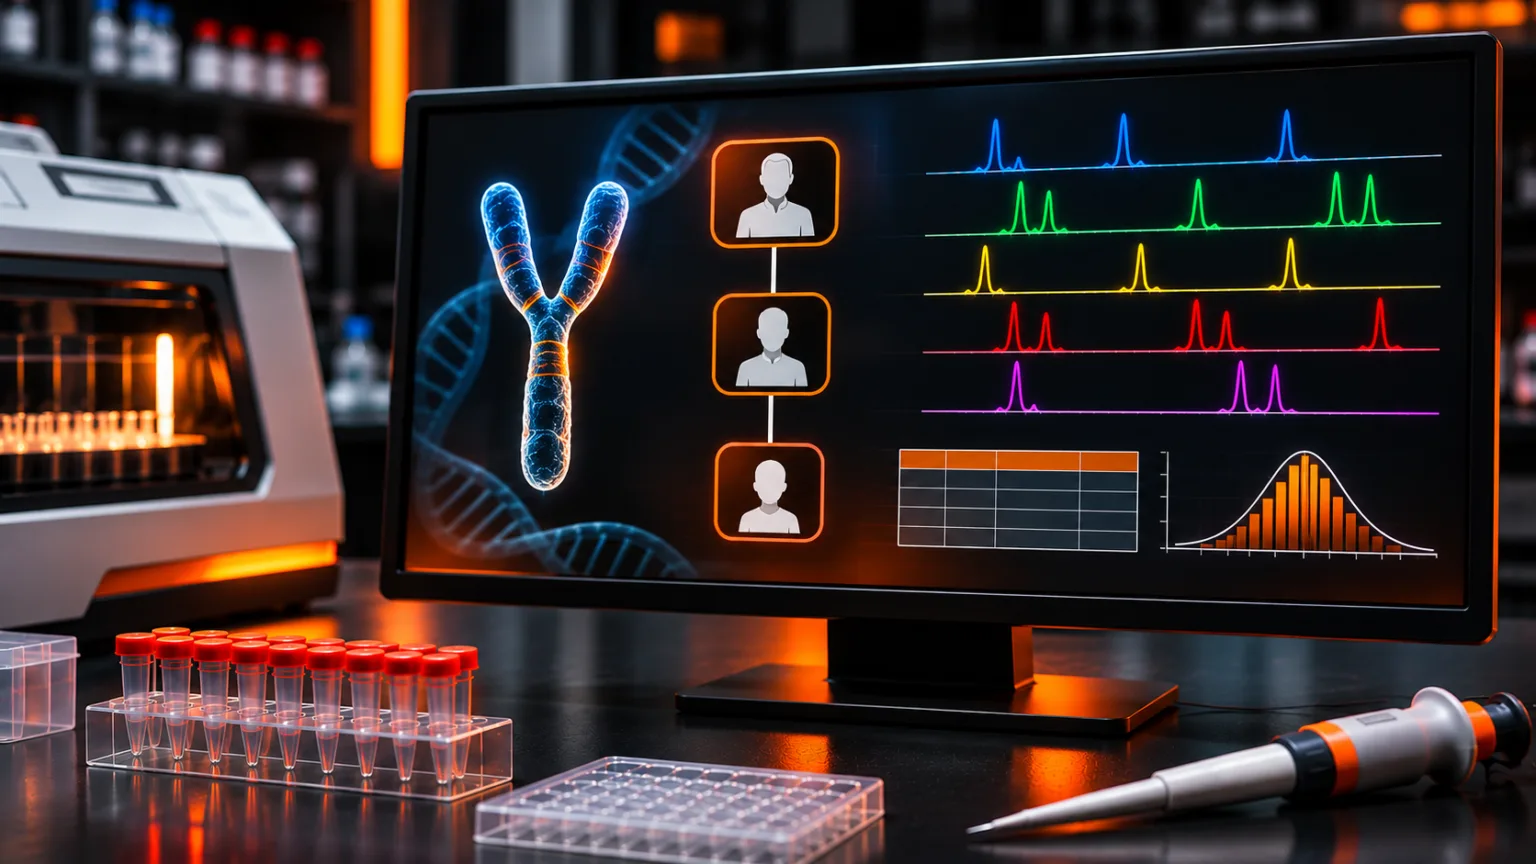
Y-chromosome short tandem repeatA short tandem repeat is a microsatellite with repeat units that are 2 to 7 base pairs in length, with the number of repeats varying among individuals, making STRs effective for human identification purposes Read Full Definition (Y-STR) profiling helps forensic laboratories detect and compare male DNA DNA, or Deoxyribonucleic Acid, is the genetic material found in cells, composed of a double helix structure. It serves as the genetic blueprint for all living organisms. Read Full Definition, particularly in samples dominated by female DNA and in investigations involving paternal relatives. Yet the interpretation is fundamentally different from an autosomal STR comparison. The Y-STR alleles
 Alleles are different forms of a gene resulting from mutations or variations in the DNA sequence or gene expression. They can be dominant (expressed with one copy) or recessive (expressed only with two copies). Read Full Definition that a man carries are transmitted together along his paternal lineage, so a matching profile can be shared by his father, brothers, sons, paternal uncles, paternal cousins or other men connected through more distant paternal ancestry. A Y-STR match is therefore not an individual identification.
- Why Y-STRs are different from autosomal STRs
- Figure 1: how a Y-STR haplotype passes through a paternal lineage
- How forensic Y-STR profiles are evaluated
- Single-source and partial haplotypes
- Artefacts, low-level peaks and multicopy loci
- Assessing the number of male contributors
- YHRD: searching for a matching Y-STR haplotype
- Observed frequency, augmented counting and confidence bounds
- Worked example 1: three matches among 1,000 haplotypes
- Worked example 2: zero YHRD matches is not zero probability
- Figure 2: where the Y-STR frequency number comes from
- Other statistical approaches: discrete Laplace, match probabilities and 2026 MatchY
- Discrete Laplace and model-based haplotype frequencies
- Why a database frequency is not automatically a case-specific match probability
- MatchY and newer pedigree-based calculations
- Population structure, metapopulations and database representation
- Paternal relatives: what does an exact haplotype match mean?
- Y-STR mutations and rapidly mutating markers
- Worked example 3: why a one-step father–son difference may occur
- Male–male Y-STR mixtures and allele sharing
- Worked example 4: an unresolved mixture with a plausible suspect haplotype
- Figure 3: why a Y-STR match is not a unique identification
- How should Y-STR results be reported?
- Checklist for analysts using YHRD and Y-STR evidence
- Frequently asked questions
- References and scientific reading
- Related Simply Forensic learning guides
The challenge is not simply finding a matching set of alleles. An analystA designated person who examines and analyzes seized drugs or related materials, or directs such examinations to be done; independently has access to unsealed evidence in order to remove samples from the evidentiary material for Read Full Definition must evaluate whether the evidence Evidence is any form of proof, such as objects, materials, or scientific findings, presented to establish or disprove a fact in a legal proceeding. It is used to reconstruct events and link or exclude individuals Read Full Definition is a reliable single-source or deduced Y-STR haplotype; determine which loci were genuinely comparable; assess the selected population database and its sampling limitations; decide how to convey the frequencyFrequency is a fundamental concept in physics and wave theory. It refers to the number of times a specific point on a wave, such as a crest or trough, passes a fixed reference point in Read Full Definition of the observed haplotype; and communicate the possibility of paternal-lineage sharing. The ISFG DNA Commission’s 2020 Y-STR recommendations emphasize that Y-STR loci are linked and that evidence weight requires specific attention to paternal relatedness, reference populations and the statistical framework.
This guide develops these ideas using **four original, fully specified hypothetical teaching examples**, a guide to YHRD searches, reporting scenarios, model comparisons, three original scientific infographics and current source-backed guidance, including the SWGDAM 2022 guidelines and 2025 supplement, the UK Forensic Science Regulator’s 2025 Y-STR guidance and peer-reviewed pedigree-based models published in 2026.
KEY LEARNING POINTS
- Y-STR loci on the non-recombining region are treated as a linked haplotype, not independently multiplied allele frequencies.
- A matching Y-STR profile is consistent with a common male source, a paternal-lineage relative or another male with the same haplotype; it is not exclusive to one man.
- Observed frequency, augmented counting, an upper confidence limit and a pedigree-based match probability are different statistical quantities.
- A YHRD search must specify the comparable loci, relevant population, sample count, search date and database release.
- Male–male mixtures, mutations and multicopy loci require explicit, validated interpretation rules before a contribution or exclusion statement is made.
Why Y-STRs are different from autosomal STRs
Most forensic autosomal STR loci lie on different chromosomes or are sufficiently separated for standard genotype-frequency calculations to employ appropriate independence assumptions (with corrections where relevant). Male-specific Y-STR markers are located on the Y chromosome A chromosome is a thread-like, rod-shaped structure found within the nucleus of a eukaryotic cell. Chromosomes are the primary carriers of genetic information in the form of genes, and they play a fundamental role in Read Full Definition and, in forensic panels, principally in its non-recombining region. A father transmits the Y chromosome to his son, with changes arising through mutation rather than the reshuffling of independently inherited alleles.
The set of reported Y-STR alleles is a haplotype. For example, a simplified profile might be DYS19 = 14, DYS390 = 24, DYS391 = 11, DYS393 = 13. Treating those four allele frequencies as independently inherited and multiplying them would be scientifically inappropriate. The forensic population-frequency question concerns the joint four-locus haplotype (and, in practice, the full validated set of comparable loci), not a product of independent locus A locus is the precise position of a gene on a chromosome. Different alleles of the same gene occupy the same locus. Read Full Definition probabilities.
Compared with autosomal typing, Y-STR analysis is valuable in sexual-assault evidence and other samples where the male contribution is weak relative to a female DNA background. The Yfiler Plus developmental-validation study describes a 27-locus assay with higher-resolution and rapidly mutating markers. Nevertheless, the ability to detect a male component does not by itself establish a man’s identity or the biological fluid deposited. See our general DNA mixture interpretation guide for mixture terminology.
| Feature | Autosomal STR | Y-STR |
|---|---|---|
| Inheritance | Typically biparental | Paternal-line transmission |
| Frequency unit | Genotype and multilocus statistical model | Linked Y-STR haplotype or lineage-aware model |
| Close relatives | Usually distinguishable with enough loci, subject to relatedness | Many paternal-line males may share the same Y haplotype |
| Female DNA background | May obscure weak male autosomal signals | Male-targeted loci can enhance detection of minor male DNA |
| Interpretation limit | Source attribution still requires appropriate statistics | Exact Y match cannot exclude paternal relatives and other male matches |
Figure 1: how a Y-STR haplotype passes through a paternal lineage
PATERNAL INHERITANCE · ORIGINAL SIMPLY FORENSIC DIAGRAM
Grandfather
Y-STR haplotype H
14 · 24 · 11 · 13
Father
Y-STR haplotype H
14 · 24 · 11 · 13
Son
H or mutational variant
14 · 24 · 11/12 · 13
An exact match can extend to paternal uncles, brothers, sons, paternal cousins and more distant paternal-line males. A mutation can introduce a difference in one transmission.
How forensic Y-STR profiles are evaluated
Single-source and partial haplotypes
At a typical single-copy Y-STR locus, a single male profile usually gives one allele. DYS385a/b and DYF387S1 are examples of multicopy marker systems in which more than one allele is expected in a single male. The analyst examines electropherograms, controls, allele designations, stutter and possible PCR artefacts before deciding which peaks constitute the interpreted haplotype.
A partial Y-STR haplotype is not automatically unusable. Its suitability depends on the loci reliably detected and the laboratory’s validationValidation, often referred to as method validation, is a crucial process in the laboratory when introducing a new machine, technology, or analytical technique. It involves a series of systematic steps and assessments to ensure that Read Full Definition. Fewer typed loci can produce more database matches and weaker discrimination. The SWGDAM 2022 guidance states that laboratories should establish the minimum locus requirements for comparison and recognize partial profiles in appropriate cases.
Artefacts, low-level peaks and multicopy loci
Y-STR electropherograms show stutter, peak-height variation, dropout, drop-in and potential artefacts much as autosomal STR profiles do. A double allele at an ordinarily single-copy Y-STR locus can arise from a male mixture, but locus duplication or another structural finding may also be relevant. A three-peak result at DYS385 does not necessarily indicate three males: the copies are not independent autosomal alleles, and the complete profile must be considered.
The stochastic thresholds and peak-height balance applicable to multicopy loci should be established empirically. In particular, a single peak at a multicopy locus does not establish that only one copy exists: allele sharing between copies or dropout can affect the observed pattern. Our STR artefact guide and low-template DNA guide give the broader analytical background.
Assessing the number of male contributors
SWGDAM recommends assessing the number of contributors based on a pattern across the profile. Multiple alleles at two or more single-copy loci generally support more than one male contributor, subject to validated interpretation and exceptions. A mixture of two men carrying identical Y-STR haplotypes might show only a single apparent haplotype, so the observed minimum number of contributors need not equal the true number. Any deduced major or minor haplotype must meet validated suitability criteria, particularly where allele sharing and stutter overlap are possible.
YHRD: searching for a matching Y-STR haplotype
The Y-Chromosome Haplotype Reference Database (YHRD) collates reference haplotypes from numerous population studies, enabling analysts to examine whether a particular haplotype has been observed and how frequently. Its database is periodically updated. Willuweit and Roewer (2015) describe the development of YHRD; the current online system must be consulted for the release applicable to a specific case rather than relying on counts reported in an old article.
Practical search workflow
- Establish the evidence haplotype: confirm allele designations, included loci, partial calls and multicopy-locus formatting against the validated interpretation.
- Choose the compatible marker set: use a haplotype resolution that is comparable across the database, documenting reductions in loci. A Yfiler Plus 27-locus case result is not automatically comparable to every lower-resolution database record.
- Identify the reference data
 Information in analog or digital form that can be transmitted or processed. Read Full Definition: select national or metapopulation subsets according to scientifically defensible ancestry and population considerations, rather than a conveniently low count.
- Record n and N: note the number of matching haplotypes n and number of reference haplotypes N in that actual search subset.
- Evaluate the statistic: distinguish raw observed frequency, augmented counting, confidence bounds and any supported model-based estimate.
- Preserve the query: record the locus set, population selection, search date, YHRD release, handling of missing/null or unusual alleles and resulting frequency method.

Databases are samples rather than a complete census of every possible paternal lineage. Collection methods, population representation, local substructure and correlated paternal relatives can influence the observed frequency. A database search with no match must be interpreted as zero observed instances in the chosen sample, not evidence that no other man on Earth carries the haplotype.
Observed frequency, augmented counting and confidence bounds
Let n be the number of matching haplotypes found among N comparable reference haplotypes. Several related but non-identical quantities arise:
| Quantity | Expression | What it communicates |
|---|---|---|
| Observed sample frequency | n / N | The observed proportion of records matching the haplotype |
| Augmented-count estimate | (n + 1) / (N + 1) | A count adjusted by a defined pseudocount convention; does not measure uncertainty by itself |
| Exact two-sided 95% binomial interval | Clopper–Pearson limits based on n successes in N trials | Uncertainty under an idealized independent binomial-sampling model |
| One-sided 95% upper confidence limit | For n = 0: 1 − 0.05^(1/N) | An upper bound with 95% one-sided nominal coverage if binomial assumptions hold |
| Two-sided 95% interval, n = 0 | Lower 0; upper 1 − 0.025^(1/N) | Different from the one-sided 95% limit because the upper tail uses 2.5% |
There is no justification for silently calling the augmented frequency a “95% confidence limit”. Nor should an estimate of the population frequency of a haplotype be automatically presented as the case-specific probability of another potential male source matching the POI. That latter question can be particularly sensitive to paternal relatedness and the way the relevant alternative-source population is defined. SWGDAM 2022 explicitly allows observed counting and augmented counting, while calling for an appropriate approach to confidence bounds and recognizing that match-probability methods are not universally settled.
Worked example 1: three matches among 1,000 haplotypes
Assume a complete, correctly typed synthetic Y-STR haplotype is searched in a relevant reference dataset with N = 1,000 independent sampled male haplotypes, and is observed n = 3 times. Treat the data as an idealized binomial sample solely for this teaching calculation. The observed proportion is 3/1,000 = 0.003 = 0.30%. The augmented-count estimate is (3 + 1)/(1,000 + 1) = 4/1,001 ≈ 0.003996 = 0.3996%.
Three different numbers for the same synthetic dataset
Observed: 3/1,000 = 0.003
Augmented: 4/1,001 ≈ 0.003996
Two-sided exact 95% binomial CI: approximately 0.000619 to 0.008742 (0.0619% to 0.8742%)
The confidence interval is calculated using the Clopper–Pearson inversion of the binomial distribution for three observed matches, not by placing a generic 95% interval around the augmented count. This example demonstrates finite-database uncertainty. Real forensic datasets can violate independent-random-sampling assumptions and may need additional caution. The count must also specify the population and exact locus panel, neither of which is provided in this synthetic example.
Worked example 2: zero YHRD matches is not zero probability
A hypothetical 27-locus haplotype is compared to an eligible dataset of N = 10,000 records, and none matches (n = 0). The observed proportion is zero. The augmented-count estimate is 1/10,001 ≈ 0.00009999, or about 0.0100%. However, the two alternative 95% confidence-bound conventions below give different results:
| Statistic | Calculation | Value |
|---|---|---|
| Augmented count | 1 / 10,001 | 0.00009999 |
| One-sided 95% upper limit | 1 − 0.05^(1/10,000) | 0.00029953 (≈ 0.02995%) |
| Two-sided exact 95% CI upper endpoint | 1 − 0.025^(1/10,000) | 0.00036882 (≈ 0.03688%) |
These percentages cannot be used interchangeably. The first is an augmented point estimate; the second is a one-sided upper confidence limit; the third is the upper endpoint of a two-sided exact 95% interval. They describe uncertainty conditional on the simple binomial sampling model, and they do not establish the number of actual men in a city who share the haplotype. **Do not invent a YHRD release or population count**: these 10,000 records are a teaching example, not a current database search.
Figure 2: where the Y-STR frequency number comes from
HAPLOTYPE SEARCH TO REPORT · ORIGINAL INFOGRAPHIC
1. Profile
Validated Y-STR haplotype and loci
2. Search
Eligible records and relevant population
3. Count
n matches / N records
4. Evaluate
Observed, augmented, CI or validated model
5. Explain
Population, assumptions, paternal sharing
Remember: YHRD estimates are conditional on the records, locus resolution, population sample and chosen statistical method. A query with zero matches is not a probability of zero.
Other statistical approaches: discrete Laplace, match probabilities and 2026 MatchY
Discrete Laplace and model-based haplotype frequencies
The discrete Laplace (DL) approach uses a statistical model of haplotype clustering and mutational relationships to estimate frequencies, including some haplotypes not observed in a database. An estimate can differ substantially from augmented counting. It is not necessarily an error when two defensible methods return different figures—they answer related questions through different assumptions. Availability depends on the marker set, model implementation and the actual population data. Do not assume that every novel, partial, multicopy or intermediate-allele haplotype can be evaluated by DL in every software configuration.
Why a database frequency is not automatically a case-specific match probability
An observed or modeled haplotype frequency describes representation in a reference population under a method. A forensic match-probability question may instead ask how likely a particular alternative man is to match a named individual, given population structure and the possibility of paternal kinship. Because many matching males belong to extended patrilines, a single “one in N” frequency can conceal important source-related dependence. Andersen and Balding (2017) argued for direct assessment of the number of matching males under population and mutation simulations. Their proposals illustrate why there is no universally accepted single summary statistic for all forensic Y-STR contexts.
MatchY and newer pedigree-based calculations
Research published in 2026 introduced MatchY, a pedigree-based approach to Y-STR match probabilities using known male pedigree relationships, mutation rates and donor-candidate status. A companion 2026 study examined factors affecting MatchY results and practical use. These models can directly address specified relationships that generic database frequencies may not capture. They are not automatically a replacement for established laboratory casework practices: any application requires defensible assumptions, suitable pedigree data, transparent propositions and proper validation.
| Approach | Principal input | Main strength | Important limitation |
|---|---|---|---|
| Observed count | n matching profiles of N | Direct description of dataset | Zero in a finite sample does not mean absence from population |
| Augmented count | (n+1)/(N+1) | Avoids a zero estimate | Point estimate alone ignores sampling uncertainty |
| Exact binomial confidence bound | n, N and chosen confidence definition | Quantifies idealized sampling uncertainty | Sampling and lineage correlations may violate model assumptions |
| Discrete Laplace | Haplotypes and population mutation/cluster model | Model-based frequencies for supported data | Model fit and marker coverage matter |
| Pedigree-based MatchY | Specified pedigree, candidate sources and mutation model | Addresses relationship-specific match questions | Requires pedigree and mutation assumptions and validation |
Population structure, metapopulations and database representation
Y-STR haplotypes can show strong population structuring because ancestral male lineages are transmitted without ordinary recombination and may expand unevenly through population history. YHRD contains different national and metapopulation groupings. The ISFG guidance explains why reference data should be chosen with reference to ancestry and relevant populations rather than modern state borders alone. The UK FSR-GUI-0013, issue 2 (March 2025) similarly discusses alternative counting methods, population selection and reporting.
For example, a small island population, a recently founded community or a patrilineally clustered group may have a Y-STR distribution that differs from a large continental reference set. A haplotype absent from a broad dataset cannot be presumed rare within every island, clan or local network. Conversely, adding extra populations without assessing their relevance is not guaranteed to improve evidential accuracyIn scientific and measurement contexts, "accuracy" refers to the degree of proximity or closeness between a measured value and the true or actual value of the measured quantity. Accuracy indicates how well a measurement reflects Read Full Definition. Laboratories should document why each population comparison was performed, identify limitations of reference coverage, and avoid selecting only the result most favourable to one side.
Y-STR haplotypes are not the same as Y-SNP haplogroups. A haplogroup describes a broader paternal phylogenetic branch defined by mutations, commonly SNPs; numerous different Y-STR haplotypes can occur within it. Predicting a haplogroup from Y-STRs is uncertain and should not be treated as proof of geographic birthplace, ethnicity, nationality or individual ancestry. Such biogeographic questions are distinct from the direct forensic question of whether an evidentiary haplotype matches a particular reference.
Paternal relatives: what does an exact haplotype match mean?
A father and son ordinarily have the same Y-STR haplotype unless a mutation changes an allele. Brothers share the father’s paternal-line chromosome subject to mutations, as do paternal uncles, male cousins connected through fathers and more distant male relatives. Distinct unrelated men can also carry a haplotype matching at a tested panel, whether through remote shared ancestry, convergence or limited marker resolution. Therefore “matching Y-STR” must never be equated with uniquely identifying a single male.
A recent simulation study, Probabilities of finding trace profile donors and their paternal relatives in Y-STR reference databases, published in Forensic Science Discover the fascinating field of Forensic Science, the application of scientific principles to legal matters. This post delves into its many disciplines, from DNA analysis to crime scene investigation, its importance in the justice system, Read Full Definition International: Genetics volume 80 (January 2026), studied matching within synthetic populations and different database fractions. In one specified 5%-sampled database scenario, match probabilities were about 41% for Yfiler Plus and 54% for PowerPlex Y23; conditional on finding a match, about 64% and 56% respectively involved a matching profile from an individual within five meioses, including the donor. These are simulation-specific results, not universal percentages for all real forensic database searches.
The evidential significance of the relative depends on the case. If a paternal brother is a plausible alternative contributor, a database estimate based on mostly unrelated males cannot simply be applied as though the brother were unrelated. Activity-level questions—such as who handled an item or when it was deposited—also need evidence beyond the Y-STR profile.
Y-STR mutations and rapidly mutating markers
Y-STRs mutate mainly by changes in the number of repeat units. The mutation rate varies substantially among loci and depends on study design and biological characteristics. A one-step difference between two males can arise over paternal transmissions; it does not automatically prove that the males are unrelated. Equally, a mismatch should not casually be dismissed as mutation when the case question is direct comparison with a crime-stain donor and validated criteria indicate a genuine exclusion. The distinction is between direct source comparison and paternal relationship analysis.
Rapidly mutating (RM) Y-STR markers such as DYS576, DYS627, DYS570 and others can improve the ability to distinguish males from the same paternal line. The Ballantyne et al. (2012) research established the utility of RM markers. More recent pedigree studies have examined their differentiation performance over many generations. Greater mutation rates bring a trade-off: they can help distinguish relatives while also producing mismatches between genuinely related men. Neither all Y-STR loci nor all RM loci have the same mutation rate.
Worked example 3: why a one-step father–son difference may occur
Consider a hypothetical four-locus father–son comparison in which the son differs by one repeat at locus A, while the other three loci match. To illustrate the basic mutation process, assume independent per-transmission mutation probabilities μ(A) = 0.020, μ(B) = μ(C) = μ(D) = 0.005. Assume every mutation at A would produce a detectable single-step change, no back mutations, and none of the other loci mutate. These values are chosen for arithmetic, not extracted from a particular kit’s empirical mutation dataset.
P(A mutates and B, C, D do not)
0.020 × 0.995 × 0.995 × 0.995 = 0.0197015 ≈ 1.970%
Under these narrowly stated assumptions, a single mutation at locus A with no changes at the other three loci has a probability of about 1.97% in one father-to-son transmission. It shows why one difference can be compatible with patrilineal transmission, but it is not a kinship likelihood ratio and it cannot be used to infer that two particular men are father and son. Real interpretation requires the correct per-locus mutation-rate models, pedigree distance, allele states, potential reversions and comparison with an alternative relationship.
By contrast, a tested crime stain whose well-resolved single-source Y-STR haplotype has several reliable discordances with a suspect’s reference would ordinarily support excluding that suspect as the direct source of that interpreted haplotype, assuming there is no plausible dropout or artefact explanation. It is a different question from whether the two males share ancestry generations earlier.
Male–male Y-STR mixtures and allele sharing
A Y-STR profile may show more than one allele at several single-copy loci because two or more males contributed. Still, the evidence may be difficult to deconvolute: two males can have the same allele at many positions, creating a single shared peak; they may have similar quantities; and stutter can overlap a genuine minor allele. Multicopy loci contribute their own baseline complexity. Accordingly, the minimum number of contributors from visible alleles is not a guarantee of the true number.
SWGDAM 2022 permits comparison of validated deduced haplotypes from distinguishable mixtures and identifies the need for laboratory support before interpreting indistinguishable mixtures. Do not simply search the union of all mixture alleles in YHRD as if it were a single-source haplotype. Nor should a YHRD frequency for a suspect’s reference haplotype be presented as a formally calculated likelihood ratio for an unresolved male–male mixture unless a validated model actually supports that inference. A general autosomal probabilistic-genotyping framework cannot be transferred to linked Y-STRs without appropriate Y-specific modelling and validation.
Worked example 4: an unresolved mixture with a plausible suspect haplotype
Suppose a synthetic evidentiary Y-STR sample shows two credible alleles at each of three single-copy loci, one allele at a fourth, and three credible alleles at a multicopy locus, after validated stutter and artefact assessment:
| Locus | Evidence alleles | Reference male S | Assessment |
|---|---|---|---|
| DYS19 (single copy) | 14, 15 | 14 | Reference allele observed |
| DYS390 (single copy) | 23, 24 | 24 | Reference allele observed |
| DYS391 (single copy) | 10, 11 | 10 | Reference allele observed |
| DYS393 (single copy) | 12 | 12 | Could be shared; does not determine NOC |
| DYS385a/b (multicopy) | 12, 14, 15 | 12, 14 | Compatible with one possible allocation, not uniquely resolved |
The two alleles at three separate single-copy loci support a mixture of at least two males in this hypothetical case. One possible explanation is that male S contributed alongside another man, with overlap at DYS393 and DYS385. Other haplotype allocations may fit the same set of peaks. Without adequate validated separation of contributors, the analyst cannot assume that every allele of S was recovered from one real contributor rather than distributed across hypothetical contributors.
One qualified educational conclusion is: “The Y-STR results indicate a mixture of DNA from at least two males. The alleles observed are compatible with the reference Y-STR haplotype of S at the evaluated positions; however, a complete individual contributor haplotype has not been reliably deduced. No database-based frequency or source attribution for S is reported for this unresolved mixture.” Any actual conclusion would depend on all typed loci, thresholds, validation and whether comparison of such an unresolved mixture is permissible under the laboratory’s SOPs.
Figure 3: why a Y-STR match is not a unique identification
THE MATCHING HAPLOTYPE CAN BELONG TO MORE THAN ONE MALE
Observed laboratory fact
The evidence and reference contain an identical interpreted Y-STR haplotype at the comparable loci.
Possible biological sources
The same male, his paternal-line relatives, or other males sharing the tested haplotype.
Inference not established
Unique identification, time of transfer, activity performed, guilt, geographic origin or ethnicity.
How should Y-STR results be reported?
For a well-resolved single-source match, the report should define the method, comparable locus set, reference population, statistic and relevant uncertainty. A suitable example is: “The Y-STR haplotype obtained from specimen Q is consistent with the Y-STR haplotype of reference male S at the loci examined. Because Y-chromosome markers are inherited along paternal lineages, this result cannot distinguish S from male relatives sharing his paternal lineage, nor from other males who happen to share the haplotype. The haplotype was observed n times among N comparable reference profiles in the stated dataset and YHRD release; the laboratory’s specified statistical estimate is reported separately.”
A report must not substitute “the DNA came from S”, “S was uniquely identified” or “only one man in 10,000 could be the source” merely because the evidence haplotype was uncommon or not found in a sample database. A profile frequency is also not the probability that S committed an offence. For general guidance, see our forensic DNA reporting guide.
| Result situation | Scientifically cautious wording | Key qualification |
|---|---|---|
| Single-source exact match | “The evidence Y-STR haplotype is consistent with the reference at the examined loci.” | Explain paternal-line sharing, unrelated matches and relevant frequency method |
| Clear single-source mismatch | “The reference is excluded as the source of the interpreted Y-STR profile.” | Confirm no mutation, allele-calling or low-template ambiguity changes the direct comparison |
| Paternal kinship mismatch | “The difference is evaluated in the context of mutation and the proposed pedigree.” | No automatic exclusion of relatedness on one-step differences |
| Unresolved male mixture | “At least two male contributors are indicated; individual haplotypes could not be resolved reliably.” | No unsupported source assignment or single-source YHRD statistic |
| No reportable Y-STR result | “No reportable Y-STR haplotype was obtained by the method used.” | Does not establish the complete absence of male DNA |
Checklist for analysts using YHRD and Y-STR evidence
- Confirm the EPG and all data-quality checks, controls, stutter and allele nomenclature.
- Document single-source/mixture status, validated NOC evaluation and any deduced component.
- Confirm comparability of kit and locus panels; handle DYS385a/b, DYF387S1 and null or unusual allele calls correctly.
- Use a scientifically justified database/metapopulation; evaluate representativeness and paternal substructure.
- Record YHRD release, search date, exact queried loci, n, N and any reductions in locus set.
- Identify the chosen estimation procedure; keep observed, augmented, interval-based and pedigree-based statistics distinct.
- Consider relatives, mutations and any alternative donor proposition actually relevant to the investigation.
- Do not convert frequency to a claim of unique identity, guilt or activity-level involvement.
- Disclose limitations of low-template data, mixtures, relatedness and population sampling.
- Apply the laboratory’s validated procedures, independent technical review and approved report wording.
Frequently asked questions
Can Y-STR DNA identify one individual?
Not uniquely on its own. The same haplotype can be shared by paternal-line males, and occasionally by other males at the tested resolution. Additional evidence is needed for individual attribution.
Why can’t Y-STR allele frequencies be multiplied?
The loci are linked on the Y chromosome and largely transmitted together; the standard independent-locus product rule for autosomal STRs does not apply. The multi-locus haplotype is treated as one linked unit or within a suitable lineage-aware model.
What does a zero-match YHRD search mean?
It means no exact matching haplotype was observed among the comparable records in the selected dataset. It does not mean the haplotype is unique or absent from the population.
What is the difference between n/N and (n+1)/(N+1)?
n/N is the observed sample proportion; (n+1)/(N+1) is an augmented-count estimate. They are not the same as a 95% confidence limit or a case-specific match probability.
Do a father and son always have identical Y-STR profiles?
They usually do, but mutations can introduce one or more differences. Interpretation of paternal relationships needs appropriate locus-specific mutation data and a defined pedigree rather than an automatic exact-match rule.
Can rapidly mutating Y-STR markers distinguish brothers?
Sometimes. RM Y-STRs increase the probability of detecting a difference between close paternal relatives, but identical profiles still occur and different loci mutate at different rates.
How do multicopy Y-STR loci affect mixture interpretation?
Some loci naturally produce more than one allele in a single male. Extra peaks there are not alone sufficient to determine the number of male contributors; laboratories must consider the whole pattern, dropout and structural variants.
Can an unresolved male mixture be searched directly in YHRD?
A composite union of mixed alleles should not be treated as one single-source haplotype. Only a suitably interpreted/deduced haplotype and compatible database query, under validated procedures, can be evaluated that way.
Is the YHRD haplotype frequency the same as an LR?
No. Frequency is an estimated representation in a database/population; an LR compares probabilities of evidence under two specified propositions. Some defined models can produce LRs, but the assumptions about relatives and competing sources matter.
What should a Y-STR match report disclose?
The comparable locus set, meaning of a match, relevant database/population and statistical method, YHRD version/search details where used, limitations and explicit paternal-lineage-sharing qualification, consistent with applicable validated SOPs.
References and scientific reading
View scientific references, standards and methods (14)
- Roewer L, Andersen MM, Ballantyne J, et al. DNA Commission of the ISFG: Recommendations on the interpretation of Y-STR results in forensic analysis. Forensic Science International: Genetics. 2020;48:102308.
- Scientific Working Group on DNA Analysis Methods (SWGDAM). Interpretation Guidelines for Y-Chromosome STR Typing by Forensic DNA Laboratories. Approved March 2, 2022.
- SWGDAM. Supplemental Information for the SWGDAM Interpretation Guidelines for Y-Chromosome STR Typing by Forensic DNA Laboratories. 2025. Official publications index.
- Forensic Science Regulator. Y-STR Profiling, FSR-GUI-0013, issue 2. England and Wales; published March 6, 2025.
- Willuweit S, Roewer L. Y chromosome haplotype reference database (YHRD): Update. Forensic Science International: Genetics. 2007;1:83–87.
- Willuweit S, Roewer L. The new Y Chromosome Haplotype Reference Database. Forensic Science International: Genetics. 2015;15:43–48.
- Y-Chromosome Haplotype Reference Database. YHRD online resource. Consult current release and search documentation at the time of casework.
- Andersen MM, Balding DJ. How convincing is a matching Y-chromosome profile? PLOS Genetics. 2017;13:e1007028.
- Ballantyne KN, Keerl V, Wollstein A, et al. A new future of forensic Y-chromosome analysis: rapidly mutating Y-STRs for differentiating male relatives and paternal lineages. Forensic Science International: Genetics. 2012;6:208–218.
- Gopinath S, Zhong C, Nguyen V, et al. Developmental validation of the Yfiler Plus PCR Amplification Kit: An enhanced Y-STR multiplex for casework and database applications. Forensic Science International: Genetics. 2016;24:164–175.
- Olsen TOS, Morling N, Eriksen PS, Kampmann ML, Mogensen HS, Andersen MM. Probabilities of finding trace profile donors and their paternal relatives in Y-STR reference databases. Forensic Science International: Genetics. Volume 80, January 2026;103320.
- MatchY research team. MatchY: A software implementation of pedigree-based calculation of Y-STR match probabilities. Forensic Science International: Genetics. 2026;84:103518.
- MatchY research team. MatchY in practice: Factors influencing pedigree-based Y-STR match probabilities. Forensic Science International: Genetics. 2026;84:103522.
- Ralf A, Montiel González D, Zandstra D, et al. Large-scale pedigree analysis highlights rapidly mutating Y-chromosomal short tandem repeats for differentiating patrilineal relatives and predicting their degrees of consanguinity. Human Genetics. 2023;142:145–160 (published online October 2022).